Ultimate First Article Approval Checklist For A New Supplier
The first commercial batch from a new intermediate supplier is not a formality. Here is the checklist, the five approval outcomes, and how to log the call.
Table of Contents
Abstract: Every assay value on a COA is measured against a reference standard — if that standard is weak, every number built on it is weak. This guide explains the primary-to-working traceability chain and the seven costly mistakes buyers make when they never ask about standards.
When a COA reports an assay of 99.6%, most buyers read it as a fact about the batch. It is not. It is the outcome of a comparison: the instrument measured how the sample responds relative to how a reference standard responds, and the calculation divided one by the other. The standard is the anchor of the entire measurement. If the standard’s assigned purity is wrong by one percent, every assay value the supplier has ever reported on that product is wrong by one percent — in the same direction.
Buyers have learned to interrogate the rest of the analytical chain. They ask whether the method was validated per ICH Q2. They read the COA line by line for red flags. They now know enough to question whether the acceptance criteria themselves are reasonable. Almost nobody asks the question underneath all of it: who calibrated your supplier’s HPLC — and how do you know that anchor is right?
This matters more for pharmaceutical intermediates than for almost any other material class, for a reason we will get to shortly: most intermediates have no pharmacopoeial standard at all. The supplier’s own qualified material is the de facto primary anchor, and the entire data package you depend on rests on how honestly and rigorously that material was characterized.
A reference standard is a fully characterized, qualified material used as the benchmark for calibration and comparison of analytical measurements. It serves three distinct functions in the laboratory, and each one puts its assigned value directly into your COA:
Regulators treat this as foundational, not technical trivia. Inspection findings on reference standards — expired standards in use, secondary standards with no qualification data, broken traceability — escalate quickly because of one logic: if the reference is weak, all QC data generated with it is weak. A buyer who adopts the same logic gets a powerful, low-effort lens on a supplier’s real quality maturity.
Reference standards are organized in a hierarchy, and each tier has a defined relationship to the one above it. The European Pharmacopoeia’s General Chapter 5.12 states the principle plainly: a secondary standard may be used for routine control purposes provided it is established with reference to the primary standard. The US framework agrees — FDA’s analytical procedures guidance describes a working standard as one “qualified against and used instead of the reference standard.”
| Tier | Source | Status | How Its Value Is Established |
|---|---|---|---|
| Primary standard | Official pharmacopoeial bodies (USP RS, Ph. Eur. CRS, Indian Pharmacopoeia) | Authoritative when cited in a monograph | Assigned by the issuing body through its own characterization program |
| Secondary standard | Established by the laboratory or an accredited reference-material producer | Acceptable for routine use only with documented comparison to the primary | Direct, replicated comparison against the primary standard using a validated method |
| Working standard | In-house material used for daily testing | Valid only within its qualification and validity period | Qualified against the primary or secondary, with an internal certificate and requalification schedule |
The economics of the chain are obvious. Primary standards are expensive and supplied in small quantities, so a laboratory that tested every batch against a pharmacopoeial vial would burn through its budget and its patience. The working standard exists to solve that: the lab qualifies a larger batch of its own material against the primary once, documents the comparison, and uses that working material for routine release testing — requalifying or replacing it on a defined schedule.
The chain is only as good as its links. Every step from primary to working adds uncertainty, and each link must remain documented: which primary lot, which comparison method, how many replicates, what acceptance criteria, what assigned value resulted. When any link is missing, the traceability that justifies the whole arrangement is gone — and the routine numbers, however precise they look, have no anchor.
Here is why this topic deserves a buyer’s attention specifically for pharmaceutical intermediates. A finished drug substance typically has a pharmacopoeial monograph and a primary standard you can buy. A custom or advanced intermediate almost never does — it does not exist in any pharmacopoeia, so there is no primary standard to purchase.
That leaves one legitimate path and one illegitimate one. The legitimate path: the supplier designates a well-characterized batch as its in-house reference standard, assigns its purity — commonly by mass balance, which is 100% minus measured organic impurities, minus water, minus residual solvents — and supports the assignment with full structural confirmation, a written qualification protocol, defined storage conditions, and periodic requalification. Done this way, an in-house standard is scientifically defensible and standard industry practice.
The illegitimate path: the supplier grabs a production batch, prints “reference standard” on the label, and uses it to release every subsequent batch — with no characterization, no assigned value, no requalification. Nothing on the COA looks different. The assay column is still populated to one decimal place. But the number is now a measurement of the batch against an unverified copy of itself, which is closer to a mirror than a standard.
The damage from a mis-assigned standard is not random noise — it is a systematic bias applied to every measurement made with it. The arithmetic makes this concrete.
Suppose a working standard’s assigned purity is 99.5%, but its true purity is 98.5%. Because the assay calculation divides the sample response by the standard response and multiplies by the standard’s purity, a 1.0% overstatement of the standard makes every sample result read approximately 1.0% higher than reality.
A batch whose true assay is 98.2% — failing a 98.0–102.0% specification only in an honest world — reports at 99.2% and passes with apparent room to spare. Multiply that across a year of batches and every marginal lot ships as conforming. Nothing in the COA looks wrong, because the error is built into the ruler, not the reading.
Laboratories that understand this compensate deliberately. A recognized practice with secondary standards is to apply a guard band — tightening internal acceptance ranges to absorb the additional uncertainty. A specification of 98.0–102.0% may be internally executed at 99.0–101.0% when the assay rests on a secondary standard, so that borderline results near the true limit do not get accepted on the strength of standard uncertainty alone. When a buyer sees a supplier whose reported results cluster suspiciously against the spec edge, one possible explanation is a standard bias quietly eating the margin.
The same logic reaches impurity numbers. Quantitation of a specific impurity depends on the response factor or reference material behind it. If the impurity standard’s assigned content is off — or if a “standard” is used that was never characterized for that impurity at all — the impurity profile on the COA is distorted in whatever direction the error points. For buyers who care about impurity control — and after reading about nitrosamine risk and elemental impurity limits, most do — the reference standard is the layer underneath the whole subject.
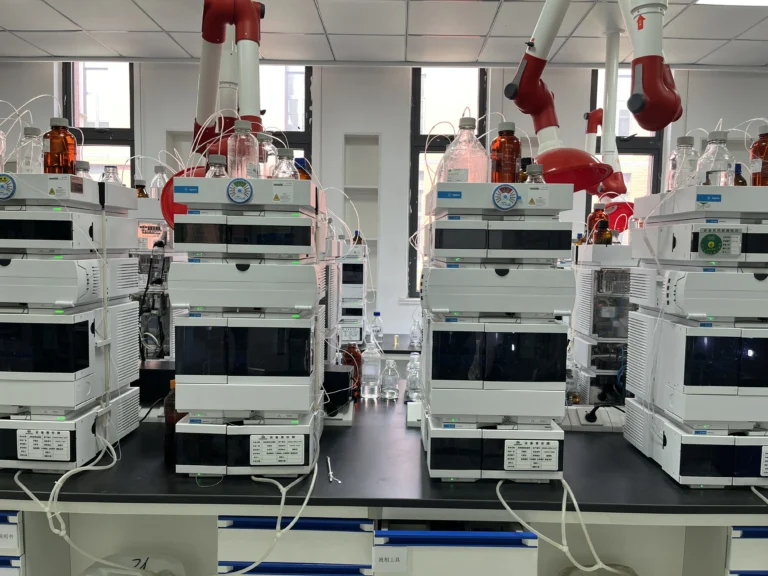
You do not need to be an analytical chemist to detect a weak standards program. These six signals are visible from the documents and answers a supplier already has, and each one warrants a follow-up question before you rely on the data.
| Red Flag | What It Means | Your Follow-Up |
|---|---|---|
| “Our reference standard is batch #XXXX of the product itself” — with no qualification data offered | Self-referential release testing; the mirror problem | Request the qualification report: characterization, assigned value, method, acceptance criteria |
| Claims of a “USP standard” for a non-compendial intermediate | No USP reference standard exists for the material — the claim is confused at best | Ask which USP catalog number; inability to answer ends the claim |
| Standard documentation shows no assigned purity value (“meets spec” is not an assigned value) | No anchor value exists to enter the assay calculation | Ask for the assigned value and how it was derived |
| No expiry or requalification date on the standard | Standards degrade too; an undated standard is an unmanaged one | Ask the requalification interval and the last execution date |
| No storage or handling conditions stated on the standard’s certificate | Light, moisture, and warmth shift purity silently | Ask how the standard is stored and monitored |
| Working standard lot changed with no bridging data between old and new | Historical results and new results no longer share one anchor | Request the bridging comparison between the two lots |
None of these questions is hostile. In fact the reaction to them is itself a screening tool — laboratories with real programs answer them from existing files, while paper-only operations deflect with urgency or vagueness. It is the same pattern you use when running a supplier audit or checking tools to verify a certificate of analysis: the document itself matters less than the system that produces it.
When you are ready to qualify or re-qualify a supplier, these are the five documents that reveal a standards program. A supplier maintaining them is doing nothing extra for you — these are working documents of a functioning quality system.
Notice what is common to all five: none of them is a new test. Every one is a record of work a serious supplier has already done. That is what makes this the most efficient diligence available to a buyer — the ask is small, and the gap between suppliers who can answer and suppliers who cannot is enormous.
Of the five documents, the internal certificate for the current working standard is the one you will look at most often, so it is worth knowing what a complete one contains. The layout varies between laboratories, but the information set does not. Here is what each field is actually telling you:
| Field | What a Complete Entry Looks Like | What It Tells You |
|---|---|---|
| Assigned value | A number with units — for example, purity 99.4% on an anhydrous basis | The anchor that enters every assay calculation; “meets specification” is not an assigned value |
| Basis of assignment | “Mass balance: 100% − 0.3% organic impurities − 0.2% water − 0.1% residual solvents” | How the number was derived — and whether each subtraction term had a real method behind it |
| Traceability statement | Qualified against primary lot / characterized in-house on date X | Which tier of the hierarchy this material belongs to, and its link upward |
| Qualification data reference | Protocol number and report number with dates | The comparison work exists in a file you can request |
| Validity / requalification date | A specific date, not “indefinite” | Whether the anchor was still valid when your batch was tested |
| Storage conditions | Specific — temperature, protection from light and moisture, desiccation | Whether the value can plausibly still be true on the day of use |
Read together, these fields answer the only question that matters: is this number anchored to something, and is the anchor still fresh? A certificate missing the assigned value or the derivation is not a thinner version of a good certificate — it is a different document entirely, because without those two fields there is nothing to trace.
One caution on interpretation: an older qualification date is not automatically bad. Standards with supporting stability monitoring can carry long validity periods. What is bad is the absence of dates, the absence of a requalification trigger list, or validity periods that never seem to end regardless of what happens to the material.
A fair objection: “I already demand ICH Q2 method validation reports. Doesn’t that cover the standard?”
No — and the reason is circularity. Method validation measures accuracy and precision relative to the standard used during validation. If the standard is biased, the method validates beautifully against the biased anchor and reports biased results with excellent precision forever. Validation establishes that the instrument and procedure compare sample to standard reliably; it cannot establish that the standard itself is true. That is a separate qualification, done separately, documented separately. The two questions — “is the method valid?” and “is the anchor true?” — must both be asked, and most buyers only ask the first.
The same layered logic applies to the numbers themselves. A COA can be arithmetically consistent, generated by a validated method, sitting inside a well-set specification — and still wrong, because the reference standard underneath was never qualified. Standards are the deepest layer of the stack: specification defines what to prove, the method proves it, the COA reports it, and the standard makes any of it mean something.
Collected in one place, these are the mistakes that cost buyers real money — usually discovered only when a downstream batch fails or an audit exposes the gap.
Reference standard qualification is the deepest layer of the quality stack, and it connects directly to the topics you are already reviewing. The specification tells the supplier what to prove about each batch. The validated method does the proving. The COA reports the results. The reference standard is what makes all of it mean something — the anchor under the entire structure.
Specialized measurements raise the stakes further. Chiral purity depends on a qualified enantiomeric reference; elemental impurity quantitation depends on traceable elemental reference solutions; and consistent results over time depend on a standard program that maintains its anchor across lot changes, which is one more reason batch-to-batch consistency starts in the standards cabinet, not on the production floor.
The practical next step is small. Add one question to your next supplier questionnaire: “Describe how the working reference standard for this intermediate is qualified, assigned, and requalified — and provide the internal certificate for the current lot.” The quality of the answer will do more sorting, in less time, than almost anything else on the form. When you want to review the standards behind a specific intermediate, contact our team.
The first commercial batch from a new intermediate supplier is not a formality. Here is the checklist, the five approval outcomes, and how to log the call.
A buyer-side decision tree for handling an intermediate supplier change notice — what to accept, contest, or reject, and how to log the call.
A supplier discontinuation is a deadline, not just bad news. The questions to ask in writing, the last-time-buy calculation, and the files to collect in time.

Manufacturer and supplier of pharmaceutical APIs and intermediates. Contact us for technical support and sourcing solutions.